ISO 14971 Risk Management Training
The design, development, and manufacturing of medical devices require a systematic approach to identifying, estimating, evaluating, controlling, and monitoring hazards. Within global regulatory frameworks, standard ISO 14971 serves as the definitive consensus standard for applying risk management principles to medical technology. Compliance with this standard is essential to satisfy the regulatory pathways of international oversight bodies, including the Therapeutic Goods Administration (TGA), the European Medicines Agency (EMA) under the Medical Device Regulation (MDR), and the United States Food and Drug Administration (FDA). Effective execution of these risk principles requires detailed knowledge of advanced risk assessment methodologies, data-driven hazard verification, and post-market signal evaluation.
Regulatory Framework and Global Harmonization of Medical Device Risk
Global medical device regulation relies on objective evidence that a device is safe and effective for its intended use. Standard ISO 14971 provides the structural architecture to evaluate safety profiles objectively. It shifts the regulatory focus from generic compliance checks to a comprehensive evaluation of the lifecycle risk-benefit profile.
Talk to QSN Academy today about ISO14971 Training
Compliance with ISO 14971 is a core prerequisite for commercialization across major international jurisdictions. In the European Union, the transition to the Medical Device Regulation (MDR 2017/745) and In Vitro Diagnostic Regulation (IVDR 2017/746) heightened the demand for rigorous risk tracking, necessitating deep alignment between risk architecture and technical documentation. In the United States, the FDA recognizes ISO 14971 as a consensus standard, making its application a vital component of Premarket Notifications [510(k)] and Premarket Approvals (PMA). This international convergence requires technical personnel to master the precise terminology, methodologies, and documentation standards defined within the framework to avoid regulatory delays.
The Systematic Risk Management Lifecycle
The application of ISO 14971 is an iterative process that spans from initial concept validation through product decommissioning. It cannot be executed as a static administrative exercise prior to product launch; it must remain an active element of the product lifecycle.
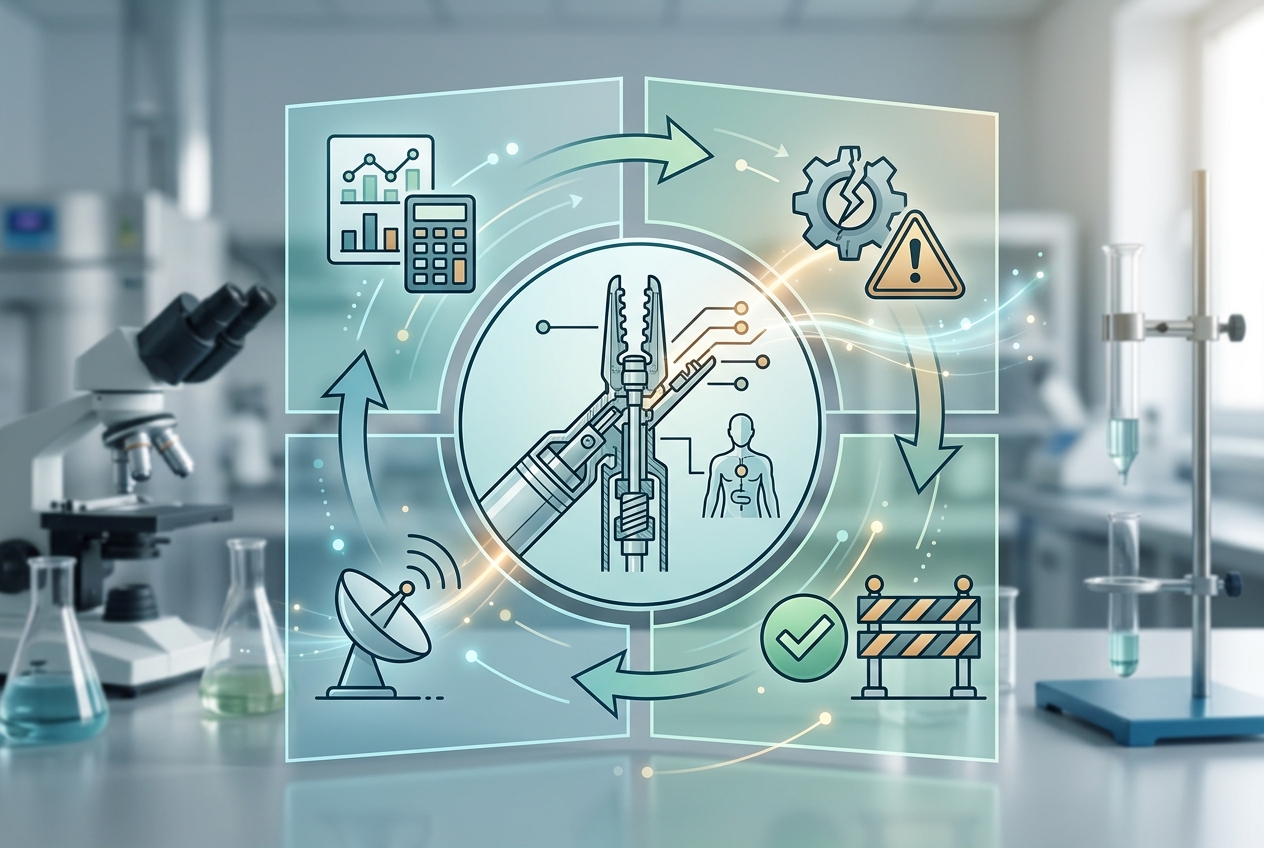
The process begins with defining the intended use and identifying characteristics related to the safety of the medical device. This step demands a rigorous engineering evaluation to establish boundaries for subsequent risk assessments. Once the operational parameters are defined, technical teams must systematically identify known and foreseeable hazards under both normal and fault conditions. These hazards typically span physical, chemical, biological, electrical, operational, and software-driven failure modes.
Risk Analysis and Risk Evaluation Metrics
Following hazard identification, personnel must perform risk analysis by estimating the risk associated with each hazardous situation. Risk is defined as the combination of the probability of occurrence of harm and the severity of that harm.
Estimating probability requires evaluating qualitative and quantitative data sources, including historical failure rates, engineering models, and scientific literature. When quantitative probability data is unavailable, particularly for novel technologies or complex software systems, teams must adopt a rigorous qualitative approach based on sequence-of-events analysis. The severity of harm is categorized using predefined criteria that range from minor discomfort to critical or fatal outcomes. The intersection of probability and severity determines whether a risk is acceptable according to the criteria defined in the risk management plan. If a calculated risk falls outside acceptable thresholds, risk control measures must be implemented systematically.
Technical Methodologies for Risk Control and Control Verification
Risk reduction must follow a strict, codified hierarchy of control measures. The standard dictates that design-inherent safety is the primary line of defense. If a hazard cannot be designed out of the device, technical teams must implement protective measures, such as physical barriers, redundant circuits, or software safety subroutines. Informational safety measures, including labeling, user alerts, and instructions for use, represent the least effective tier of control and are only applied when design and protective options are exhausted.
Once risk controls are selected, engineering teams must verify their implementation and effectiveness through formal verification and validation activities. This involves compiling objective data to prove that the control successfully reduces the probability or severity of harm without introducing new, unforeseen hazards. The residual risk remaining after control implementation is then evaluated to confirm it conforms to the organization's overarching risk acceptability criteria.
Advanced Analytical Tools: FMEA versus FTA
Accurate risk estimation relies on selecting appropriate analytical tools suited to the technical complexity of the device. Two primary methodologies widely utilized within compliant risk management systems are Failure Mode and Effects Analysis (FMEA) and Fault Tree Analysis (FTA).
FMEA is a bottom-up, inductive analytical technique that evaluates individual component failures and traces their effects upward through the system architecture to determine the ultimate impact on the end-user. This approach is highly effective for mechanical, electrical, and hardware sub-assemblies where individual component failure rates can be quantified reliably. Conversely, FTA is a top-down, deductive analytical methodology that begins with an undesired system-level event (the "top event") and works downward to identify the combination of component failures, human errors, and environmental factors that could cause it. FTA is especially valuable for complex systems, software integration, and combination products where multiple concurrent faults must occur to create a hazardous situation. Competency in both methodologies allows organizations to construct comprehensive risk profiles that address both single-fault and multi-fault scenarios.
Post-Market Surveillance and Signal Detection
The risk management process does not conclude when a medical device receives market authorization. ISO 14971 places significant emphasis on the collection and evaluation of production and post-production information. This feedback loop ensures that the initial assumptions made during the design phase remain valid when exposed to real-world clinical use.
Organizations must establish systematic mechanisms to gather data from production processes, clinical literature, user complaints, regulatory adverse event databases, and service records. This information must be analyzed periodically to detect signals indicating that the frequency or severity of a hazard exceeds initial estimations, or that entirely new hazards have emerged. When post-market data reveals a deviation from the established risk profile, technical teams must re-evaluate the risk management file, initiate CAPA protocols, and update the risk-benefit analysis to maintain regulatory alignment and ensure patient safety.
Structuring the Risk Management File for Regulatory Audits
The ultimate output of the risk management process is the Risk Management File (RMF). The RMF serves as the definitive compilation of objective evidence demonstrating that the manufacturer has systematically managed risk in accordance with ISO 14971.
A compliant RMF must contain the risk management plan, hazard identifications, risk estimations, control verifications, and the final risk management report signed by authorized technical personnel. During regulatory audits and notified body assessments, the RMF is subjected to intense scrutiny. Reviewers assess the logical traceability from identified hazards to risk control verification data. Fragmented documentation, unsubstantiated probability assertions, and failure to update the RMF with post-market data represent common non-conformances that can jeopardize market access. Specialized training programs provided by QSN Academy equip technical personnel with the structural methodologies needed to build, maintain, and audit an airtight Risk Management File that stands up to international regulatory inspections.
